
The US FDA has posted warning letters to three Chinese active pharmaceutical ingredient (API) makers over failed records requests. Two refused to hand over records and a third gave an incomplete response. The letters are dated 29 October and 3 November 2026; FDA's pages carry the date 10 November. No Irish or EU firm was named.
All three letters rest on section 704(a)(4) of the US Federal Food, Drug, and Cosmetic Act, which lets FDA demand records "in advance of or in lieu of" an inspection. None of the three letters describes an FDA inspection of the firm that received it. The cases show FDA using paperwork alone to block imports, a route that applies to any API plant that ships to the US, including plants in Ireland.
- Who: Chongqing Qiancan Technology Co., Ltd. and Tianjin Healthcare Medical Technology Co., Ltd. (both letters dated 29 October 2026), and Nanjing Hanxin Pharmaceutical Technology Co. Ltd. (3 November 2026).
- Why: Qiancan and Tianjin Healthcare refused to provide requested records. Nanjing Hanxin's response left out an API it had registered.
- Trigger: FDA import records showed APIs for US pharmacy compounding naming Qiancan and Tianjin Healthcare as manufacturer.
- Consequence: all three firms' drugs are already on FDA import alerts, which allow detention at the US border without physical examination.
- Ireland: no Irish or EU company is named in these letters. The lesson for exporters is about records readiness.
What did FDA say to each firm?
| Firm and letter | What FDA found |
|---|---|
| Chongqing Qiancan Technology Co., Ltd., Chongqing; letter 320-27-14, 29 October 2026 | Records requested on 17 June 2026. The firm de-registered on 29 June, replied on 30 June that it could not meet FDA requirements, and re-registered on 13 July. Import Alert 66-79 from 26 August 2026. |
| Tianjin Healthcare Medical Technology Co., Ltd., Tianjin; letter 320-27-15, 29 October 2026 | Records requested on 17 June 2026. On 24 June the firm said it would stop making the products for the US and declined to send records; it declined again after an FDA follow-up on 25 June. Import Alert 66-79 from 25 August 2026. |
| Nanjing Hanxin Pharmaceutical Technology Co. Ltd., Nanjing; letter 320-27-19, 3 November 2026 | Records requested on 18 November 2025. The response did not document manufacture of an API active under its registration. The firm later said production had moved to a subsidiary, whose drugs FDA had placed on an import alert after an inspection. Import Alert 66-40 from 22 June 2026. |
Why does refusing a records request matter so much?
For Qiancan and Tianjin Healthcare, the refusal itself is the violation. Both letters cite section 301(e) of the Act, which makes refusing access to required records a prohibited act. Both say: "Your firm refused to provide the duly requested records." FDA adds that without them it has no indication of the level of quality assurance for drugs listed as made at the facilities.
Tianjin Healthcare argued it had stopped supplying the US. FDA rejected that, noting the imported APIs were still within expiry. Ceasing distribution, the letter says, "does not retroactively relieve you of your regulatory obligations". FDA also reminded the firm that 21 CFR 207.29 requires registration to be updated within 30 days of ceasing operations.
Both firms were placed on Import Alert 66-79, "Detention Without Physical Examination of Drugs From Foreign Establishments Refusing FDA Inspection". FDA's text for that alert covers firms that delay, deny or limit an inspection, and names a 704(a)(4) records request as one form of assessment.
What went wrong at Nanjing Hanxin?
Nanjing Hanxin did respond, but FDA found a gap. Its registration listed an API that its records did not cover. When FDA asked again, the firm said production had moved to a subsidiary after November 2025. That subsidiary had also been qualified as the firm's supplier of a critical starting material, and FDA had already found its drugs adulterated after an inspection. The letter states: "Your oversight of your suppliers is deficient."
FDA called the API adulterated under section 501(a)(2)(B) because the quality unit failed to ensure CGMP compliance. It raised concern that naming Nanjing Hanxin instead of the actual manufacturer may have been "an attempt to circumvent safeguards". The firm must give FDA a supplier qualification plan, stability data supporting retest dates for US-market APIs, a three-year list of drugs shipped with their real manufacturers, and written notices to affected customers.
What should EU and Irish API exporters check?
None of these letters involves an Irish or EU firm. The same legal tool applies to every foreign plant registered with FDA, and FDA's final guidance "Conducting Remote Regulatory Assessments: Questions and Answers" (June 2025) also covers section 704(a)(4) requests. Points drawn from the three letters:
- Watch the registration inbox. FDA sent both June requests to the email contact in the firms' drug registrations. That address needs an owner and a response procedure.
- Keep registration and listing accurate. Every API listed should be covered by records the site can produce. De-registering after a request did not help Qiancan.
- Be able to produce batch and quality records quickly. For APIs, FDA's CGMP expectations are set out in its ICH Q7 guidance, and EU GMP Part II is based on the same ICH Q7 text. Both treat documentation, batch records and laboratory records as core requirements. Finished-dose plants work under 21 CFR Parts 210 and 211.
- Remember material still in the market. FDA said records remain required while imported APIs are within expiry, even after US sales stop.
- Qualify and oversee suppliers. If a related company makes a starting material or the API itself, the records must show who made what.
- Hold stability data behind retest dates. FDA asked Nanjing Hanxin for exactly this.
Our guide to FDA warning letters and Irish sites explains how letters work, and our earlier report on the FDA warning letter to Sanofi's Waterford plant covers an inspection-based case.
Frequently asked questions
What is an FDA 704(a)(4) records request?
It is a legal demand for records or other information that FDA can make before an inspection or instead of one. Compliance is mandatory, and refusing access to required records is a prohibited act.
Were any Irish companies named?
No. The three letters went to firms in Chongqing, Tianjin and Nanjing, China.
What does an import alert do?
It lets US border officials detain a firm's shipments without physically examining them. A firm can ask FDA for removal, and the letters say Qiancan and Tianjin Healthcare may request an FDA inspection.
Why does compounding come up?
FDA found the APIs from Qiancan and Tianjin Healthcare in shipments intended for US pharmacy compounding, which is how the firms came to its attention.
How long do the firms have to reply?
Each letter asks for a written response within 15 business days of receipt.
Outlook
FDA says it may withhold approval of new applications or supplements that list these firms until they comply, and may keep refusing their shipments. Priya Life Science will report any FDA action involving an Irish site.
Sources
- US FDA, Warning letter 320-27-14 to Chongqing Qiancan Technology Co., Ltd., 29 October 2026
- US FDA, Warning letter 320-27-15 to Tianjin Healthcare Medical Technology Co., Ltd., 29 October 2026
- US FDA, Warning letter 320-27-19 to Nanjing Hanxin Pharmaceutical Technology Co. Ltd., 3 November 2026
- US FDA, Import Alert 66-79 and Import Alert 66-40
- Federal Register, Conducting Remote Regulatory Assessments: Questions and Answers; Guidance for Industry; Availability, 26 June 2025
- US FDA, Q7 Good Manufacturing Practice Guidance for Active Pharmaceutical Ingredients, September 2016
- European Commission, EudraLex Volume 4, Part II: Basic Requirements for Active Substances used as Starting Materials
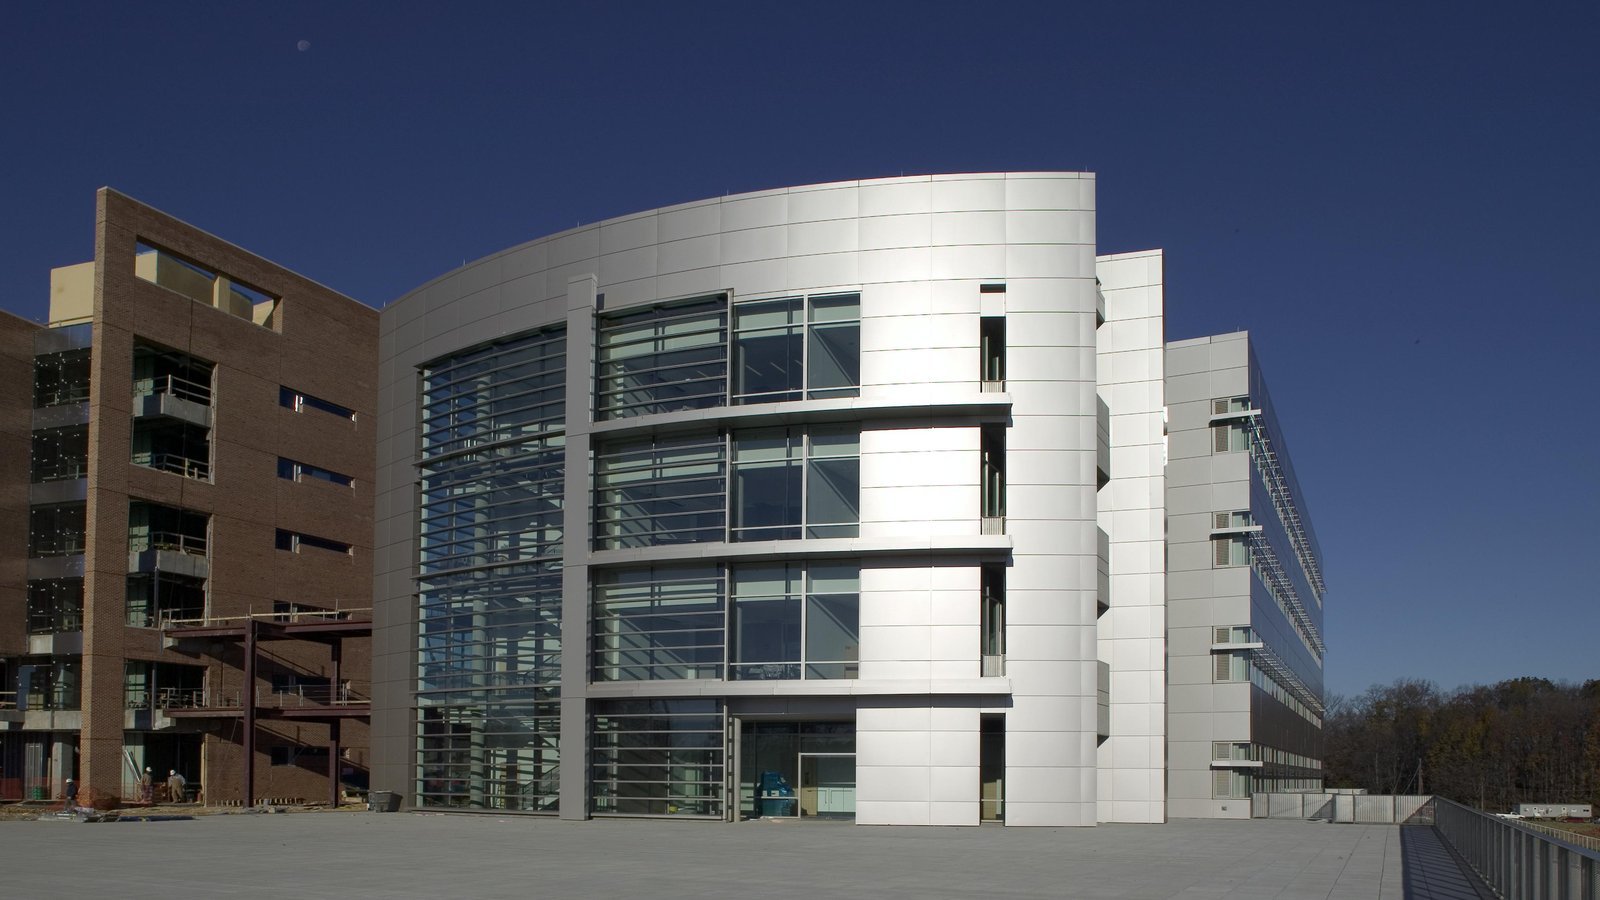
Cover image: Main entrance of FDA Building 1 at the agency's White Oak campus, Silver Spring, Maryland. Photo: U.S. Food and Drug Administration, public domain, via Wikimedia Commons, cropped.